
2.5: Solubility in an aqueous world - The Hydrophobic Effect
- Page ID
- 69576
\( \newcommand{\vecs}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\( \newcommand{\vecd}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash {#1}}} \)
\( \newcommand{\dsum}{\displaystyle\sum\limits} \)
\( \newcommand{\dint}{\displaystyle\int\limits} \)
\( \newcommand{\dlim}{\displaystyle\lim\limits} \)
\( \newcommand{\id}{\mathrm{id}}\) \( \newcommand{\Span}{\mathrm{span}}\)
( \newcommand{\kernel}{\mathrm{null}\,}\) \( \newcommand{\range}{\mathrm{range}\,}\)
\( \newcommand{\RealPart}{\mathrm{Re}}\) \( \newcommand{\ImaginaryPart}{\mathrm{Im}}\)
\( \newcommand{\Argument}{\mathrm{Arg}}\) \( \newcommand{\norm}[1]{\| #1 \|}\)
\( \newcommand{\inner}[2]{\langle #1, #2 \rangle}\)
\( \newcommand{\Span}{\mathrm{span}}\)
\( \newcommand{\id}{\mathrm{id}}\)
\( \newcommand{\Span}{\mathrm{span}}\)
\( \newcommand{\kernel}{\mathrm{null}\,}\)
\( \newcommand{\range}{\mathrm{range}\,}\)
\( \newcommand{\RealPart}{\mathrm{Re}}\)
\( \newcommand{\ImaginaryPart}{\mathrm{Im}}\)
\( \newcommand{\Argument}{\mathrm{Arg}}\)
\( \newcommand{\norm}[1]{\| #1 \|}\)
\( \newcommand{\inner}[2]{\langle #1, #2 \rangle}\)
\( \newcommand{\Span}{\mathrm{span}}\) \( \newcommand{\AA}{\unicode[.8,0]{x212B}}\)
\( \newcommand{\vectorA}[1]{\vec{#1}} % arrow\)
\( \newcommand{\vectorAt}[1]{\vec{\text{#1}}} % arrow\)
\( \newcommand{\vectorB}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\( \newcommand{\vectorC}[1]{\textbf{#1}} \)
\( \newcommand{\vectorD}[1]{\overrightarrow{#1}} \)
\( \newcommand{\vectorDt}[1]{\overrightarrow{\text{#1}}} \)
\( \newcommand{\vectE}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash{\mathbf {#1}}}} \)
\( \newcommand{\vecs}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\(\newcommand{\longvect}{\overrightarrow}\)
\( \newcommand{\vecd}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash {#1}}} \)
\(\newcommand{\avec}{\mathbf a}\) \(\newcommand{\bvec}{\mathbf b}\) \(\newcommand{\cvec}{\mathbf c}\) \(\newcommand{\dvec}{\mathbf d}\) \(\newcommand{\dtil}{\widetilde{\mathbf d}}\) \(\newcommand{\evec}{\mathbf e}\) \(\newcommand{\fvec}{\mathbf f}\) \(\newcommand{\nvec}{\mathbf n}\) \(\newcommand{\pvec}{\mathbf p}\) \(\newcommand{\qvec}{\mathbf q}\) \(\newcommand{\svec}{\mathbf s}\) \(\newcommand{\tvec}{\mathbf t}\) \(\newcommand{\uvec}{\mathbf u}\) \(\newcommand{\vvec}{\mathbf v}\) \(\newcommand{\wvec}{\mathbf w}\) \(\newcommand{\xvec}{\mathbf x}\) \(\newcommand{\yvec}{\mathbf y}\) \(\newcommand{\zvec}{\mathbf z}\) \(\newcommand{\rvec}{\mathbf r}\) \(\newcommand{\mvec}{\mathbf m}\) \(\newcommand{\zerovec}{\mathbf 0}\) \(\newcommand{\onevec}{\mathbf 1}\) \(\newcommand{\real}{\mathbb R}\) \(\newcommand{\twovec}[2]{\left[\begin{array}{r}#1 \\ #2 \end{array}\right]}\) \(\newcommand{\ctwovec}[2]{\left[\begin{array}{c}#1 \\ #2 \end{array}\right]}\) \(\newcommand{\threevec}[3]{\left[\begin{array}{r}#1 \\ #2 \\ #3 \end{array}\right]}\) \(\newcommand{\cthreevec}[3]{\left[\begin{array}{c}#1 \\ #2 \\ #3 \end{array}\right]}\) \(\newcommand{\fourvec}[4]{\left[\begin{array}{r}#1 \\ #2 \\ #3 \\ #4 \end{array}\right]}\) \(\newcommand{\cfourvec}[4]{\left[\begin{array}{c}#1 \\ #2 \\ #3 \\ #4 \end{array}\right]}\) \(\newcommand{\fivevec}[5]{\left[\begin{array}{r}#1 \\ #2 \\ #3 \\ #4 \\ #5 \\ \end{array}\right]}\) \(\newcommand{\cfivevec}[5]{\left[\begin{array}{c}#1 \\ #2 \\ #3 \\ #4 \\ #5 \\ \end{array}\right]}\) \(\newcommand{\mattwo}[4]{\left[\begin{array}{rr}#1 \amp #2 \\ #3 \amp #4 \\ \end{array}\right]}\) \(\newcommand{\laspan}[1]{\text{Span}\{#1\}}\) \(\newcommand{\bcal}{\cal B}\) \(\newcommand{\ccal}{\cal C}\) \(\newcommand{\scal}{\cal S}\) \(\newcommand{\wcal}{\cal W}\) \(\newcommand{\ecal}{\cal E}\) \(\newcommand{\coords}[2]{\left\{#1\right\}_{#2}}\) \(\newcommand{\gray}[1]{\color{gray}{#1}}\) \(\newcommand{\lgray}[1]{\color{lightgray}{#1}}\) \(\newcommand{\rank}{\operatorname{rank}}\) \(\newcommand{\row}{\text{Row}}\) \(\newcommand{\col}{\text{Col}}\) \(\renewcommand{\row}{\text{Row}}\) \(\newcommand{\nul}{\text{Nul}}\) \(\newcommand{\var}{\text{Var}}\) \(\newcommand{\corr}{\text{corr}}\) \(\newcommand{\len}[1]{\left|#1\right|}\) \(\newcommand{\bbar}{\overline{\bvec}}\) \(\newcommand{\bhat}{\widehat{\bvec}}\) \(\newcommand{\bperp}{\bvec^\perp}\) \(\newcommand{\xhat}{\widehat{\xvec}}\) \(\newcommand{\vhat}{\widehat{\vvec}}\) \(\newcommand{\uhat}{\widehat{\uvec}}\) \(\newcommand{\what}{\widehat{\wvec}}\) \(\newcommand{\Sighat}{\widehat{\Sigma}}\) \(\newcommand{\lt}{<}\) \(\newcommand{\gt}{>}\) \(\newcommand{\amp}{&}\) \(\definecolor{fillinmathshade}{gray}{0.9}\)(Learning goals written by Claude, Sonnet 4.6, Anthropic)
Free Energy, Chemical Potential, and Partition Equilibria
- Define chemical potential (μ) as the incremental change in free energy per mole of a substance, apply the equation G = G° + RT ln[A] to describe solute distribution between compartments, and derive the condition for equilibrium in terms of equal chemical potentials across phases.
- Write the equilibrium expression for a solute partitioning between aqueous and organic phases, relate the partition coefficient Kpart to ΔG° for transfer, and explain why Kpart values are pharmacologically important for predicting membrane permeability of drugs.
Thermodynamics of the Hydrophobic Effect
- Interpret experimental thermodynamic transfer data (ΔG°, ΔH°, ΔS°) for the movement of aliphatic alcohols from pure liquid to water, and explain the counterintuitive finding that transfer of nonpolar groups into water is enthalpically favorable but entropically disfavored — identifying the entropic cost as the dominant contribution to the hydrophobic effect.
- Explain the molecular basis of the unfavorable entropy of hydration for nonpolar solutes: the formation of a structured, "ice-like" water cage around the solute reduces the number of accessible microstates for solvent water, and dissolution of this cage upon transfer of the solute back to the nonpolar phase restores positional entropy to the bulk solvent.
- Interpret the linear relationship between Δμ° and hydrocarbon chain length (approximately −25 cal/Ų) as evidence that each added methylene group makes a constant, additive contribution to the free energy of transfer, and explain what this linearity implies about the molecular-level additivity of hydrophobic interactions.
- Distinguish the hydrophobic effect from simple hydrophobic interactions: the former is a thermodynamic driving force rooted in solvent entropy gain upon release of ordered water, while the latter refers to the induced dipole-induced dipole attractive interactions between nonpolar groups themselves.
Introduction
Many biomolecules, such as triacylglycerols, cholesterol esters, and waxes, are nonpolar. Other biomolecules, such as proteins and many lipids, have both polar and nonpolar parts. We know from experience that oil floats on water, indicating that it is less dense than water and doesn't dissolve in it. You have also probably performed liquid-liquid extractions in chemistry labs, using the solubility properties of nonpolar molecules to extract them from a water-based mixture and transfer them to a more nonpolar phase, such as octanol or chloroform. To understand the stability of biomolecules with nonpolar parts in aqueous solutions, we need to consider not only their noncovalent interactions with water (which we explored in Chapter 2.4) but also the thermodynamics of their interactions in aqueous environments.
We have been taught and internalized the notion that "like dissolves like." We anthropomorphize molecules, saying that nonpolar molecules "like" nonpolar environments. We can rationalize solubility properties by examining a molecule's noncovalent attractive and repulsive interactions in an aqueous solution. So we usually focus on enthalpic contributions to stability. What about entropy? We should consider net changes in noncovalent solute-solute, solute-solvent, and solvent-solvent interactions, along with their thermodynamic contributions to overall stability. To consider the thermodynamics of solubility in water, we need to determine the ΔG (free energy change) for all processes involved.
The Change in Free Energy (G) and Chemical Potential (μ)
ΔG, the free energy change for a reaction, determines the spontaneity and extent of a chemical or physical reaction. A system's free energy depends on three variables: temperature T, pressure P, and n, the number of moles of each substance. For the latter, consider solute X on two sides of a permeable membrane. If the concentration of X is the same on each side, as shown in Figure \(\PageIndex{1}\), the system is in equilibrium.

If the system is composed of two different parts, A and B, the system is at equilibrium (ΔG=0) if TA = TB, PA = PB, and the change in the absolute free energy per mole of A is ΔGA/Δn = ΔGB/Δn. More precisely, using simple calculus, we would discuss incremental changes in absolute free energy/mol, dGA/dn for A, which is the chemical potential of A (μA), and dGB/dn (μB) for B. At equilibrium, dGA/dn = dGB/dn. We will use the free energy (G) here, but μ later in this section. G is the absolute free energy/mol (again, chemical potential), where G=Go +RTln[A]. You can derive the following equation from the equations you used in introductory chemistry (which we reviewed in Chapter 1.3).
\begin{equation}
\begin{array}{l}
\Delta \mathrm{G}=\Delta \mathrm{G}^{0}+\mathrm{RTIn} \mathrm{Qr} \\
\Delta \mathrm{G}=\Delta \mathrm{H}-\mathrm{T} \Delta \mathrm{S} \\
\Delta \mathrm{G}^{0}=\Delta \mathrm{H}^{0}-\mathrm{T} \Delta \mathrm{S}^{0} \\
\Delta \mathrm{G}^{0}=-\mathrm{RTInK} \mathrm{eq}
\end{array}
\end{equation}
Now, let's apply this to the chemical equation for the solubility of a given solute in water. You eventually reach a saturation point if you add a sparingly soluble hydrocarbon (HC) or sodium chloride to water. The salt solution is saturated with dissolved NaCl, and no further increase in NaCl (aq) occurs. The solution reaches saturation for a sparingly soluble hydrocarbon, after which phase separation occurs.
Let's add a drop of a slightly soluble hydrocarbon liquid (HCL) to water, as shown in the diagram below. At t=0, the system is not at equilibrium, and some of the HC will transfer from the pure liquid to water, so at time t=0, ΔGTOT < 0. This is illustrated in Figure \(\PageIndex{2}\).

The following equations can be derived.
\begin{equation}
\begin{array}{c}
\Delta \mathrm{G}_{\mathrm{TOT}}=\left(G_{\mathrm{HC}-\mathrm{W}}\right)-\left(G_{\mathrm{HC}-\mathrm{L}}\right)=\mathrm{G}_{\mathrm{HC}-\mathrm{W}}^{0}+R T \ln [\mathrm{HC}]_{\mathrm{W}}-\left(\mathrm{G}_{\mathrm{HC}-\mathrm{L}}^{0}+R T \ln [\mathrm{HC}]_{\mathrm{L}}\right)= \\
\Delta \mathrm{G}_{\mathrm{TOT}}=\left(\mathrm{G}_{\mathrm{HC}-\mathrm{W}}^{0}-\mathrm{G}_{\mathrm{HC}-\mathrm{L}}^{0}\right)+R T \ln \left([\mathrm{HC}]_{\mathrm{W}}-\ln [\mathrm{HC}]_{\mathrm{L}}\right)= \\
\Delta \mathrm{G}_{\mathrm{TOT}}=\Delta \mathrm{G}^{0}+R T \ln \frac{[\mathrm{HC}]_{\mathrm{W}}}{[\mathrm{HC}]_{\mathrm{L}}}
\end{array}
\end{equation}
Now, add a bit more complexity to the last example. Add a hydrocarbon x to a biphasic system of water and octanol as shown in Figure \(\PageIndex{3}\). Shake it vigorously. At equilibrium, x would have "partitioned" between the two mostly immiscible phases.

A simple reaction can be written for this system: X aq ↔ X oct.
If X is a hydrocarbon, ΔG < 0 for the reaction written above. Also, ΔGo < 0, since this term is independent of concentration and depends only on the intrinsic stability of X in water compared to that of octanol. This simple equation holds:
\begin{equation}
\Delta \mathrm{G}_{\mathrm{TOT}}=\left(\mathrm{G}_{\mathrm{X}-\mathrm{oct}}^{0}-\mathrm{G}_{\mathrm{X}-\mathrm{w}}^{0}\right)+R T \ln \frac{[\mathrm{X}]_{\mathrm{oct}}}{[\mathrm{X}]_{\mathrm{w}}}=\Delta \mathrm{G}^{0}+R T \ln \frac{[\mathrm{X}]_{\mathrm{oct}}}{[\mathrm{X}]_{\mathrm{w}}}
\end{equation}
At equilibrium, ΔG=0 and the equation can be rewritten as:
\begin{equation}
\Delta \mathrm{G}^{0}=-R T \ln \frac{[\mathrm{X}]_{\mathrm{oct}}}{[\mathrm{X}]_{\mathrm{w}}}=-\mathrm{RTlnK}_{\mathrm{part}}
\end{equation}
Kpart is the equilibrium partition coefficient for X in octanol and water. You can readily determine this value in the lab. Just shake a separatory flask with a biphasic system of octanol and water after injecting a bit of X. Then separate the layers and determine the concentration of X in each phase. Plug these numbers into the last equation. You should be able to predict the sign and relative magnitude of ΔGo since it depends only on the intrinsic stability of the molecules in the different environments, not on concentration. Kpart values are often determined for drugs because they must diffuse across cell membranes to enter the cytoplasm, where they can act. Drugs, hence, must have a reasonable Kpart to pass through the membrane but not so high that they are insoluble.
Introduction to the Hydrophobic Effect
Now let's ask this question: What are the enthalpic and entropic contributions to the ΔG for interacting a nonpolar hydrocarbon (HC) with water? For this section, we will replace ΔG with Δμ (the change in chemical potential, but we will use these terms interchangeably. Likewise, we will use this equation: Δμo = ΔHo - T ΔSo.
Also, instead of framing the reaction as the dissolution of an organic molecule in water, we will frame it as the transfer of a hydrocarbon X from an aqueous solution to the pure hydrocarbon liquid (HC) or
\begin{equation}
\mathrm{X}(\mathrm{aq}) \leftrightarrow \mathrm{X}(\mathrm{HC})
\end{equation}
Figure \(\PageIndex{4}\) shows the standard free energies of transfer of a hydrocarbon X from an aqueous solution to a pure liquid hydrocarbon (HC), X (aq) ↔ X (HC). where
\begin{equation}
\Delta \mu^{\circ}=\mu^{\circ} x(H C)-\mu^{\circ} x(a q)
\end{equation}
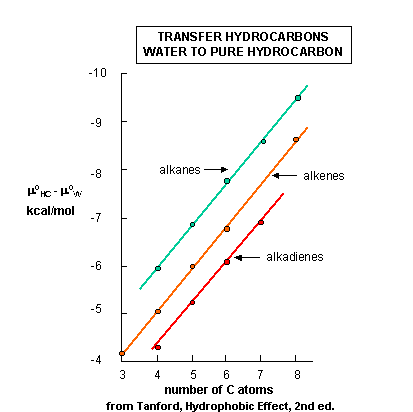
Δμo is negative because a transfer back to the pure HC is favored on stability grounds. In each graph, Δμo is less than 0, and the value of Δμo decreases (gets more negative as you go up the y-axis, which shows increasingly negative values of Δμo) in a linear fashion with increasing numbers of carbon atoms in the alkyl chain. Notice how straight and parallel the lines are. Nature is speaking to us in these figures. By determining the surface area of the hydrocarbon molecules and the decrease in Δμo with each added CH2 (methylene group), one can calculate that the Δμo decreases by 25 cal/Å2 (105 J/Å2) per methylene added.
We expected that Δμo for transferring X from a pure liquid HC would be negative. We could get more information if we could determine both the entropic and enthalpic contributions. Such data are presented in the table below, which shows the transfer of short, single-chain alcohol X (an amphiphile with a polar head and a longer nonpolar "tail") from the pure liquid alcohol (ROH) to water (the opposite of the previous figures)
\begin{equation}
X(R O H) \leftrightarrow X(W)
\end{equation}
Thermodynamic Parameters for Transfer of Aliphatic Alcohol X from the Pure Liquid to Water at 25 °C (enthalpy determined by calorimetry)
| alcohol X | μw0-μ ROH0 kcal/mol (kJ/mol) | Hw0-H ROH0 kcal/mol (kJ/mol) | Sw0-S ROH0 cal/deg mol (J/deg mol) |
(Cp)w0-(Cp)ROH0 cal/deg mol |
| ethanol | 0.760 (3.18) | -2.43 (-10.2) | -10.7 (-44.8) | 39 (163) |
| n-propanol | 1.58 (6.61) | -2.42 (-10.2) | -13.4 (-56.1) | 56 (234) |
| n-butanol | 2.4 (10) | -2.25 (-9.41) | -15.6 (-65.3) | 72 (301) |
| n-pentanol (solubility 22g/L H2O) |
3.22 (13.5) | -1.87 (-7.82) | -17.1 (-71.5) | 84 (352) |
We expect Δμo to become increasingly positive as the chain length increases and water solubility becomes increasingly disfavored. What is perplexing about this data is not that the transfer of these ROHs to water is disfavored, but that the transfer is enthalpically favored (negative ΔH0). This seems counterintuitive since it goes against the adage that "like dissolves like," as discussed earlier. From an enthalpic point of view, the amphiphiles prefer (albeit marginally) to be in water. Entropy disfavors this reaction. The data show that the nonpolar molecule "prefers" not to be in water because it is entropically disfavored.
At first glance, you might guess that the entropy should favor the movement of ROHs into the water since they could access a larger volume and have greater freedom of motion. Therefore, ROHs in water have more possible microstates. However, this is only part of the process. We haven't considered water's entropy. A literal cavity must be created to accommodate a hydrocarbon in water. The creation of this more ordered cavity must be entropically disfavored (again because the process proceeds to a state with fewer microstates and lower positional entropy).
In the reverse process, transferring the hydrocarbon from water to the pure liquid dissipates the cavity, thereby increasing the number of available microstates for the released solvent, bulk water. This entropic contribution favors moving a hydrocarbon from water to the pure hydrocarbon lipid. This "hydrophobic effect" is the main thermodynamic driving force for moving organic molecules out of water.
Imagine this scenario. When you place a hydrocarbon group into water, water seeks (admittedly an anthropomorphic term) to maintain its hydrogen bonding. Hence, it is forced into a more ordered structure around the HC to maintain its H-bonding, characterized by fewer microstates. We will explore the hydrophobic effect in greater detail in a future chapter.
How can we explain the favorable enthalpic contribution of placing a nonpolar molecule into water? Again, this goes against our adage: "like dissolves like." The negative ΔH suggests interactions among all the participants are more favorable when the nonpolar group is in water. One source of such interactions could be the highly structured water in the "cage" surrounding the nonpolar molecule. If it were more structured than bulk water—hence more "ice-like"—then forming these extra H-bonds would contribute to the negative enthalpy change. When the nonpolar molecule is removed from the water, which proceeds with a positive ΔH, the "ice-like" water cage would "melt"; like ice melting, this is not favored enthalpically, as heat must be added. Heat energy must be supplied to break the H-bonds as ice melts into liquid water. This molecular model for understanding the thermodynamic data may be simplistic, but for now, let's use it.
Summary
(Summary written by Claude, Sonnet 4.6, Anthropic)
This chapter extends the thermodynamic framework developed in earlier sections to the specific and biologically critical problem of how nonpolar molecules behave in aqueous environments. The central concept—the hydrophobic effect—is far more subtle than the familiar heuristic "like dissolves like" suggests, and understanding it requires carefully separating enthalpic and entropic contributions to the free energy of transfer.
The chapter opens by formalizing the concept of chemical potential (μ), the partial molar free energy that governs how a substance distributes itself between compartments or phases. At equilibrium, the chemical potential of a solute must be equal in all phases it can access; any imbalance drives net transfer until equality is restored. For a solute X partitioning between an aqueous phase and an organic phase such as octanol, the equilibrium is described by a partition coefficient Kpart, which is directly related to ΔG° for transfer. Since ΔG° reflects only the intrinsic relative stability of the solute in each environment — independent of concentration — its sign and magnitude can be predicted from structural principles. Partition coefficients have direct pharmacological relevance: a drug must be sufficiently nonpolar to partition into and diffuse across lipid membranes, yet sufficiently polar to remain soluble enough in the aqueous phase to reach its target.
Thermodynamic analysis of hydrocarbon transfer from a pure liquid organic phase into water yields a result that initially seems to contradict chemical intuition. Experimental calorimetric data for short-chain aliphatic alcohols show that transfer into water is actually enthalpically favorable (ΔH° < 0) — water forms slightly stronger interactions with the amphiphile than the amphiphile forms with itself in its pure liquid state, possibly through the formation of a structured, "ice-like" hydrogen-bonded cage of water molecules around the nonpolar portion of the solute. The transfer is thermodynamically disfavored overall because of entropy: ΔS° is negative and becomes more negative with chain length, reflecting the reduction in accessible microstates as water molecules are forced into an ordered, clathrate-like cage around the buried nonpolar group. Conversely, the reverse process — transfer of the nonpolar group from water back into a pure hydrocarbon phase — is entropically driven by the release of these ordered water molecules into the bulk, restoring their translational and rotational freedom. This increase in solvent entropy upon collapse or burial of nonpolar surfaces is the thermodynamic signature of the hydrophobic effect.
A particularly striking feature of the transfer free energy data is its linearity with chain length: Δμ° decreases by approximately 25 cal/Ų for each additional nonpolar methylene group. This remarkable regularity — visible in straight, parallel lines across alkane, alkene, and alkyne series — indicates that each methylene unit makes an independent, additive, and constant contribution to the driving force for transfer out of water. This proportionality between hydrophobic free energy and buried surface area becomes a quantitative tool used throughout structural biology and drug design to estimate the thermodynamic cost or benefit of exposing or burying nonpolar surfaces in proteins, membranes, and ligand-binding interfaces. The hydrophobic effect, properly understood as an entropy-driven expulsion of nonpolar groups from water rather than a simple attraction between nonpolar molecules, is one of the most important organizing principles in biochemistry.