
22.1: Overview of Nitrogen Metabolism - Nitrogenase
- Page ID
- 15178
\( \newcommand{\vecs}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\( \newcommand{\vecd}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash {#1}}} \)
\( \newcommand{\dsum}{\displaystyle\sum\limits} \)
\( \newcommand{\dint}{\displaystyle\int\limits} \)
\( \newcommand{\dlim}{\displaystyle\lim\limits} \)
\( \newcommand{\id}{\mathrm{id}}\) \( \newcommand{\Span}{\mathrm{span}}\)
( \newcommand{\kernel}{\mathrm{null}\,}\) \( \newcommand{\range}{\mathrm{range}\,}\)
\( \newcommand{\RealPart}{\mathrm{Re}}\) \( \newcommand{\ImaginaryPart}{\mathrm{Im}}\)
\( \newcommand{\Argument}{\mathrm{Arg}}\) \( \newcommand{\norm}[1]{\| #1 \|}\)
\( \newcommand{\inner}[2]{\langle #1, #2 \rangle}\)
\( \newcommand{\Span}{\mathrm{span}}\)
\( \newcommand{\id}{\mathrm{id}}\)
\( \newcommand{\Span}{\mathrm{span}}\)
\( \newcommand{\kernel}{\mathrm{null}\,}\)
\( \newcommand{\range}{\mathrm{range}\,}\)
\( \newcommand{\RealPart}{\mathrm{Re}}\)
\( \newcommand{\ImaginaryPart}{\mathrm{Im}}\)
\( \newcommand{\Argument}{\mathrm{Arg}}\)
\( \newcommand{\norm}[1]{\| #1 \|}\)
\( \newcommand{\inner}[2]{\langle #1, #2 \rangle}\)
\( \newcommand{\Span}{\mathrm{span}}\) \( \newcommand{\AA}{\unicode[.8,0]{x212B}}\)
\( \newcommand{\vectorA}[1]{\vec{#1}} % arrow\)
\( \newcommand{\vectorAt}[1]{\vec{\text{#1}}} % arrow\)
\( \newcommand{\vectorB}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\( \newcommand{\vectorC}[1]{\textbf{#1}} \)
\( \newcommand{\vectorD}[1]{\overrightarrow{#1}} \)
\( \newcommand{\vectorDt}[1]{\overrightarrow{\text{#1}}} \)
\( \newcommand{\vectE}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash{\mathbf {#1}}}} \)
\( \newcommand{\vecs}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\(\newcommand{\longvect}{\overrightarrow}\)
\( \newcommand{\vecd}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash {#1}}} \)
\(\newcommand{\ket}[1]{\left| #1 \right>}\)
\(\newcommand{\bra}[1]{\left< #1 \right|}\)
\(\newcommand{\braket}[2]{\left< #1 \vphantom{#2} \right| \left. #2 \vphantom{#1} \right>}\)
\(\newcommand{\braopket}[3]{\left< #1 \vphantom{#2}\vphantom{#3} \right| #2 \vphantom{#1}\vphantom{#3} \left| #3 \vphantom{#1}\vphantom{#2} \right>}\)
\(\newcommand{\qmvec}[1]{\mathbf{\vec{#1}}}\)
\(\newcommand{\op}[1]{\hat{\mathbf{#1}}}\)
\(\newcommand{\expect}[1]{\langle #1 \rangle}\)
\(\newcommand{\dfn}[1]{\emph{\textbf{#1}}}\)
(Learning goals written by Claude, Sonnet 4.6, Anthropic)
The Nitrogen Cycle and Thermodynamics of Fixation
- Describe the major transformations of the biological nitrogen cycle — fixation (N₂ → NH₄⁺), nitrification (NH₄⁺ → NO₂⁻ → NO₃⁻), denitrification (NO₃⁻ → N₂), anammox (NO₂⁻ + NH₄⁺ → N₂), and ammonification — identifying the oxidation state of nitrogen in each species, whether each reaction is aerobic or anaerobic, and why human perturbation through Haber-Bosch-derived fertilizers has overwhelmed the cycle's natural balancing capacity.
- Explain why N₂ reduction to NH₃ is thermodynamically favorable at room temperature (ΔG° = −16.4 kJ/mol, exothermic ΔH° = −46.2 kJ/mol) yet kinetically inert — connecting the high activation energy to the N≡N triple bond's stability — and describe how the Haber-Bosch process resolves this kinetic barrier using high pressure (to shift equilibrium toward fewer gas molecules), elevated temperature (to overcome activation energy despite reducing Keq), and a magnetite catalyst.
Structure and Metal Cofactors of Nitrogenase
- Describe the two-component architecture of nitrogenase — the Fe-protein reductase homodimer (containing the 4Fe-4S F cluster, ferredoxin/flavodoxin binding sites, and two ATP-binding sites) and the MoFe-protein catalytic heterodimer (containing the 8Fe-7S P cluster between α and β subunits and the FeMo-cofactor M cluster in the α subunit where N₂ reduction occurs) — explaining the directional flow of electrons from the F cluster through the P cluster to the M cluster and how two ATPs are hydrolyzed per electron transferred to drive subunit dissociation and reassociation.
- Describe the structural features of the FeMo-cofactor M cluster — including its Fe₇MoS₉ core, interstitial carbide derived from SAM's methyl group, molybdenum coordination to three sulfur ions and 3-hydroxy-3-carboxy-adipic acid, and the role of His195 (as probable general acid/base for the first N₂ protonation) and Val70 (as a gatekeeper controlling substrate access) — and explain why nitrogenase is irreversibly inactivated by O₂, making symbiosis with anaerobic microenvironments essential for nitrogen-fixing organisms.
Mechanism of N₂ Reduction: The Lowe-Thorneley Model and Organometallic Reactions
- Describe the Lowe-Thorneley model of nitrogenase catalysis — tracing the sequential addition of four electrons and four protons (E₀ → E₄) to the FeMo-cofactor, explaining how two electrons reduce two protons to form bridging hydrides (Fe-H-Fe, three-center two-electron bonds) rather than reducing N₂, and why the E₄ "Janus intermediate" is the obligate entry point for N₂ binding, with the overall stoichiometry requiring 16 ATP and 8 electrons (6 for N₂ reduction, 2 for obligate H₂ production).
- Explain the organometallic reactions central to nitrogenase mechanism — including oxidative addition (H₂ insertion into the reduced Fe metal center increases Fe oxidation state, producing bridging hydrides), reductive elimination (the coupled reverse process releases H₂ as the Fe center is reduced, creating the activated low-valent 4Fe face that binds N₂ in a kinetically inert push-pull mechanism), and migratory insertion (transfer of hydride from Fe to N during reduction of diazene N₂H₂ and subsequent intermediates) — and explain why bridging rather than terminal hydrides are critical for storing reducing equivalents without premature loss as H₂ upon protonation.
- Trace the second half of the N₂ reduction mechanism — describing how N₂ binds to the activated E₄ state and is immediately converted to diazene (HN=NH) as H₂ is released by reductive elimination (using two H⁺ from sulfide-bound protons and the two bridging hydrides as reducing equivalents), followed by sequential migratory insertions of hydrides into N-H bonds to produce successively diazenido, hydrazido, and ultimately two NH₃ molecules — and explain how this mechanism maintains Fe oxidation state changes of no more than ±1 unit throughout the catalytic cycle.
Introduction
Organic chemistry is usually described as the chemistry of carbon-containing molecules. But isn't that definition a bit carbon-centric, especially given the prevalence of oxygen-containing molecules? What about nitrogen? We live in a dinitrogen-rich atmosphere (80%), and all classes of biomolecules (lipids, carbohydrates, nucleic acids, and proteins) contain nitrogen. Dinitrogen is very stable due to its triple bond and nonpolar nature. We rely on a few organisms to fix N2 from the atmosphere to form ammonium (NH4+), which, through nitrification and denitrification, can form nitrite (NO2-), nitrate (NO3-), nitric oxide (NO), and nitrous oxide (N2O), the latter being a potent greenhouse gas. We'll concentrate on the metabolic fate of amino groups in amino acids and proteins in the next section. Before exploring their fates, look at Figure \(\PageIndex{1}\), which shows an overall view of the biological nitrogen cycle. The study of biochemistry should encompass more than homo sapiens and expand to the ecosystem in which we are such a small but damaging part.

Figure \(\PageIndex{1}\): Nitrogen Cycle
Let's break down the diagram from a biochemical perspective. There are aerobic and anaerobic processes (conducted by bacteria). Nitrogen-containing substances include both inorganic (ammonium, nitrate, nitrite) and organic (amino acids, nucleotides, etc.) molecules. The reactions shown are both oxidative and reductive (note: the oxidation numbers of the nitrogen atoms in the molecules are shown in red). Most reactions occur underground and are carried out by bacterial and Archaeal microorganisms.
Here are some of the major reactions:
- N2 fixation (a reduction): Soil prokaryotes use the enzyme nitrogenase to reduce N2 from the air to ammonium (NH4+). The energetically unfavorable reaction requires a lot of ATP. Primary producers, such as plants, can then take up the ammonium and incorporate it into biomolecules, such as amino acids, which animals consume. For those who may still believe that people have marginal effects on our biosphere, consider this. We may soon fix more N2 to NH3 through the industrial Haber-Bosch reaction (used for fertilizer and explosive production), which is all made by the biosphere. Much of the nitrogen in use comes from the Haber-Bosch reaction. The excess NH4+ (upwards of 50%) produced industrially and entering the soil through fertilizers (mostly as NH4NO3) has overwhelmed nature's ability to balance the nitrogen cycle and is not taken up by plants. Microorganisms metabolize it into nitrite and nitrate.
- Nitrification: Ammonia-oxidizing aerobic microorganisms convert ammonium to nitrite, and a separate group of nitrite-oxidizing aerobic bacteria converts it further to nitrate. Here are the reactions (Rx 1 and 2) to produce nitrate through a hydroxylamine intermediate, followed by the formation of nitrate (Rx 3).
NH3 + O2 + 2e- → NH2OH + H2O Rx 1
NH2OH + H2O → NO2- + 5H+ + 4e- Rx 2
NO2- + 1/2 O2 → NO3- Rx 3
These added ions exceed soil capacity and end up in runoff water, polluting our rivers and lakes.
- Denitrification: This anaerobic reaction pathway converts nitrate into N2. Here is the net reaction:
2NO3- + 10e- + 12H+ → 2N2 + 6H2O
- Anammox reaction: This recently discovered bacterial anaerobic reaction pathway converts ammonium and nitrate to nitrogen gas (N2). Here is the net reaction
NO2- + NH3+ → N2 + 2H2O
- Ammonification (not to be confused with mummification) occurs when plants and animals decompose, releasing ammonium back into the soil for reuse by plants and microbes.
These reactions are shown in the abbreviated Nitrogen Cycle shown in Figure \(\PageIndex{2}\).

Figure \(\PageIndex{2}\): Abbreviated Nitrogen Cycle
Nitrogen metabolites are essential nutrients for plants, playing a crucial role in regulating plant growth (primary productivity) and maintaining biodiversity in the biosphere. All living organisms require feedstocks to produce energy and as substrates for biosynthetic reactions. What is used depends on the organism. Plants are primary producers, so they use their synthesized carbohydrates for both energy production and biosynthesis. For carnivores, proteins and their derived amino acids serve as an energy source through oxidation and as biosynthetic precursors for various cellular processes. In omnivores, the energy source depends on the "fed" state. With abundant food resources, carbohydrates and lipids serve as the primary energy sources. Unlike carbohydrates and lipids, which can be stored as glycogen and triacylglycerols for later use, excess protein and its amino acids cannot be stored, so amino acids are eliminated or used for oxidative energy.
In the fed state, carbohydrates are the main source, while in the unfed state, lipids take a predominant role. Under starvation conditions, the organism breaks down its own proteins and uses them for oxidative energy production and continued biosynthesis. In diseased states like diabetes, which can be likened to starvation despite abundant carbohydrates, both lipids and amino acids become energy sources.
How are amino acids oxidatively metabolized in animals? Several pathways can achieve this. Still, it seems logical that NH4+ would be removed, and the carbons in the remaining molecule would eventually enter glycolysis or the TCA cycle in the form of ketoacids. NH4+ is toxic in high concentrations. In humans, ammonium is not oxidized to nitrite or nitrate, unlike in soil, where microorganisms facilitate oxidation. The body can recycle it back into nucleotides or amino acids, and it eliminates excess amounts. Both processes must be highly controlled. We will turn our attention to the oxidation of amino acids in the next section.
Nitrogenase: An Introduction
Beauty is in the eye of the beholder.
As the domain of biochemistry covers the entire biological world, the extent of coverage of a given topic in textbooks can depend, in part, on the interest and experiences of the author(s) presenting the material. Should relevance determine coverage? If so, books focused on human or medical biochemistry would surely omit photosynthesis. If topics are selected based on their importance for life, then photosynthesis must surely be covered. If so, then nitrogenase must also be included. Given the chemical difficulty of the reaction and the remarkable elegance of the evolved biochemical solution, both photosynthesis and nitrogen fixation must be presented. Although nitrogen fixation is a reductive reaction, it shares strong similarities with the oxygen-evolving complex in photosynthesis. Both catalyze enormously important redox reactions involving an abundant atmospheric gas, using a complex, unique inorganic metallic cofactor that evolution has selected as uniquely suited for the task.
Every first-year chemistry student can draw the Lewis structure of dinitrogen, N2, which contains a triple bond and a lone pair on each nitrogen. If Lewis structures resonate with them, they should be able to state that the triple bond makes N2 extraordinarily stable, thus explaining why we can breathe an atmosphere containing 80% N2 without dying. If they have taken biology, they are also aware that very few biological organisms can utilize N2 as a substrate, as this requires breaking bonds between the nitrogen atoms, a chemical process reserved for nitrogen-fixing bacteria found in rhizomes of certain plants. Lastly, they likely memorized that the Haber-Bosch process requires high pressure and temperature to convert N2 and H2 into ammonia (NH3). As with any scientific advance, the Haber-Bosch process has brought both harm (it is used for explosive weapons) and benefit (in the form of fertilizers). This process now fixes enough N2 in fertilizers to support half of the world’s population, with nitrogenase supporting the rest. Efforts are underway to genetically modify plants to produce nitrogenase, eliminating the need for fertilizers, but this may create unforeseen problems.
You might be surprised to learn that at room temperature, the equilibrium constant favors ammonia formation, so ΔG0 < 0. The reaction is favored enthalpically as it is exothermic at room temperature. It is disfavored entropically, as should be evident from the balanced equation:
\[\ce{N2(g) + 3H2(g) → 2 NH3(g)}. \nonumber \]
The thermodynamic parameters for the reaction (per mol) are ΔH° = –46.2 kJ, ΔS° = –389 J K–1, and ΔG° = –16.4 kJ at 298 K
Entropy is negative because the reaction proceeds from 4 to 2 molecules. From an enthalpy perspective, raising the temperature of an exothermic reaction drives it in the reverse direction. Increasing the pressure shifts the equilibrium toward the side with the fewest molecules.
If the reaction is favored thermodynamically at room temperature, why doesn’t it proceed? This story sounds familiar, as this same descriptor applies to the oxidation of organic molecules with dioxygen. There, we showed using MO theory that the reaction is kinetically slow. Same with NH3 formation. Superficially, we must break bonds in stable N2 to initiate the reaction, resulting in a high activation energy and sluggish kinetics.
One could jump-start the reaction by raising the temperature, but that would slow an exothermic reaction. The Keq (or KD) and ΔG0 depend on temperature, and for this reaction, the reaction becomes disfavored at higher temperatures. Bosch's solution was high pressure, which shifts the reaction to the side with fewer gas molecules, and high temperature, which overcomes the activation energy barrier and makes the reaction kinetically feasible. A complex metal catalyst (magnetite, Fe3O4 -with metal oxides like CaO and Al2O3) prevents reduction of Fe with H2, provides an absorptive surface to bring reagents together, and facilitates bond breaking in H2 and N2.
In photosynthesis, the oxygen-evolving complex (OEC), which contains Mn, Fe, S, and Ca, oxidizes another highly stable and ubiquitous molecule, H2O. Now we explore the remarkable mechanisms of the nitrogenase complex, which reduces N2 to NH3.
What might be needed to drive this reaction biologically? You might surmise the list to include:
- a source of energy, most likely ATP, to facilitate this complex reaction;
- a source of electrons as the N atoms move from an oxidation state of 0 in elemental N to 3- in NH3; this source turns out to be a protein called flavodoxin or ferredoxin. Of course, these electrons also have an interesting origin before they are in the electron carriers of these proteins;
- some pretty amazing metal centers accept and donate electrons in a controlled way; these centers are mostly FeS clusters with an additional cluster containing molybdenum (Mo). The clusters are named F, P, and M.
- a source of hydrogen; you might have guessed correctly that it’s not H2 gas (from where would that come?), but H+ ions, which are pretty ubiquitously available.
- a net reaction that is different from the Haber-Bosch process (N2 + 3H2 → 2 NH3).
Here is the actual reaction catalyzed by nitrogenase:
\[\ce{N2 + 8e^{-} + 16ATP + 8H^{+} → 2NH3 + H2 + 16ADP + 16P_i}. \nonumber \]
Let’s think a bit about the reaction. As electrons are added, the attraction between the nitrogen atoms must decrease. Eventually, the bonds between them must break. Protons can be added easily to maintain charge neutrality. A basic mechanism might involve intermediates as shown in Figure \(\PageIndex{3}\).
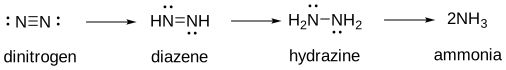
Figure \(\PageIndex{3}\): Possible intermediates in the conversion of dinitrogen to ammonia by nitrogenase
Nitrogenase can also interact with other small molecules with triple bonds, including C≡O: and H-C≡C-H.
The Structure of Nitrogenase
Nitrogenase is a multiprotein complex in which the functional biological unit is built from two sets of the following dimeric structures:
- a homodimer of subunits, E and F, which have binding sites for the mobile carrier of electrons (the protein ferredoxin or flavodoxin), ATP, and an FeS cofactor (4Fe-4S, called the F cluster) that accepts electrons. These subunits are therefore called the nitrogenase reductase subunits.
- a heterodimer of alpha and beta subunits. The a (alpha chain) binds the 8Fe-7S F cluster and the Fe-S-Mo M cluster. These subunits comprise the (di)nitrogenase catalytic subunits. The iron-molybdenum M cluster is in the α subunit and is where N2 reduction occurs. The P-cluster is between the α and β subunits and facilitates electron flow between the Fe-protein (F cluster) and FeMo-cofactor (M cluster)
For clarity, one-half of the overall structure of the protein complex with bound ATP and metal centers is shown in Figure \(\PageIndex{4}\).
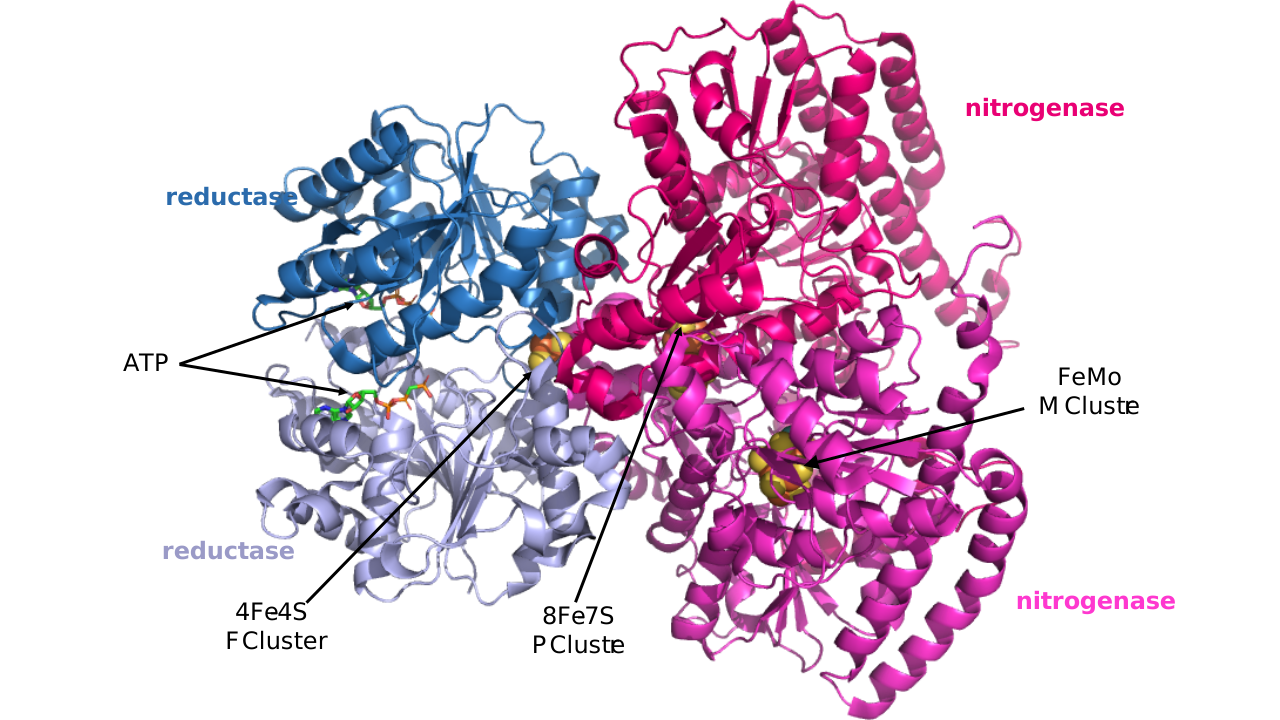
Figure \(\PageIndex{4}\): Nitrogenase structure (4wzb)
This half-structure consists of a homodimer of the reductase monomers and a heterodimer of nitrogenase subunits.
Figure \(\PageIndex{5}\) shows an interactive iCn3D model of the half-structure of the nitrogenase complex from Azotobacter vinelandii (4WZB) (long load).

 Figure \(\PageIndex{5}\): Nitrogenase complex from Azotobacter vinelandii (4WZB). (Copyright; author via source). Click the image for a popup or use this external link: https://structure.ncbi.nlm.nih.gov/i...zPNjfPRwq8MSeA (long load))
The structure is color-coded similarly to Figure 4. The F cluster is labeled as SF4, the P cluster as CLF (FE(8)-S(7) cluster), and the M cluster as ICS (iron-sulfur-molybdenum cluster with interstitial carbon).
The reductase subunits (called Av2 in Azotobacter vinelandii), accept electrons from ferredoxin and is where the ATP analog, ACP (phosphomethylphosphonic acid adenylate ester), and the 4Fe-4S F cluster is bound. The nitrogenase subunits (called Av1 in Azotobacter vinelandii) convert N2 to NH3, and it is where the 8Fe-7S P cluster, FeMo (C Fe7 Mo S9) M Cluster is bound.
An enhanced view of the bound cofactors and ATP is shown in the same spatial orientation in Figure \(\PageIndex{6}\)below.

Figure \(\PageIndex{6}\): Bound cofactor metal clusters and ATP in nitrogenase
You can easily imagine the direction of electron flow from the F cluster to the P cluster to the M cluster.
The metal centers are shown in Figure \(\PageIndex{7}\) in more detail, both in line and spacefill views.
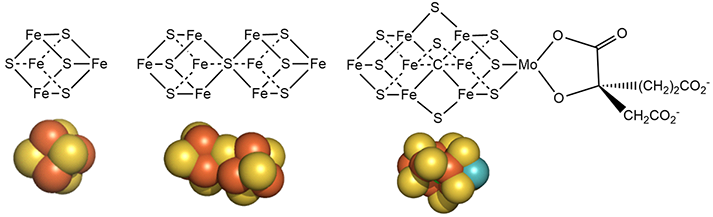
Figure \(\PageIndex{7}\): Detailed structures of the metal cofactors in nitrogenase
Mo is bound to 3 sulfur ions and oxygen from 3-hydroxy-3-carboxy-adipic acid as shown in Figure 7 above.
The M cluster contains an interstitial carbide ion derived from the -CH3 group attached to the sulfur of S-adenosyl-methionine (SAM), allowing the carbide to be labeled with either 13C or 14C for mechanistic studies. These labeled carbides are neither exchanged nor used as a substrate during the enzyme's catalytic turnover. Thus, the carbide likely stabilizes the M cluster. The figures below show more detailed mechanisms but without the carbide.
We will focus on two key features of the reaction mechanism: ATP hydrolysis and the flow of electrons and protons to N2 as it is reduced to NH3.
O2 destroys the enzyme. Plants are aerobic. So how does it function? See Chapter 32.17: Part 4 - Fixing Nitrogen Fixation for details!
ATP hydrolysis
ATP binds in the reductase subunit (AV2) where ferredoxin brings in electrons, and where the F metal cluster is bound. As GTP hydrolysis controls conformational change and subunit dissociation in the heterotrimeric Gαβγ in signal transduction, ATP hydrolysis in the reductase (AV2) subunits drives not only electron transfer but also dissociation/reassociation of the reductase and nitrogenase catalytic subunits. It appears that two ATPs are hydrolyzed per electron transferred from the F cluster to the M cluster. Since the oxidation numbers of N are 0 and -3 in N2 and NH3, respectively, sequential rounds of ATP hydrolysis and dissociation/reassociation occur. The Fe-protein hydrolyzes ATP only when bound to the MoFe-protein.
The overall reaction reduces N2 to two molecules of NH3 at the FeMo cofactor. It involves the reductive elimination of hydrides bridged by Fe ions (Fe-H-Fe) in a reaction that also produces H2 as a by-product. We'll describe this complex reaction next.
Nitrogenase Reaction: Part 1 - Addition of Electrons and Protons
The sequential path of electrons from the reductase subunit containing the F cluster to the P and M clusters in the nitrogenase subunit should be apparent from the figures above. We will focus on N2 binding and how it receives electrons from the M cluster. Figure \(\PageIndex{8}\) shows the FeMo-cofactor and some adjacent amino acid residues. Mo is labeled but not shown in spacefill. HCA is a bound molecule of 3-hydroxy-3-carboxy-adipic acid, which interacts with Mo. The carbide is shown in the middle as the green sphere and labeled CX.
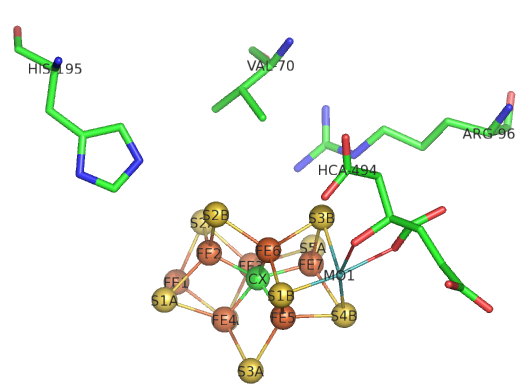
Figure \(\PageIndex{8}\): Side chains near the M cluster
If Val 70 is mutated to Ile, a substrate appears not to access the cluster, suggesting that N2 may interact with the top part of the structure, with the residues shown acting as gatekeepers. His side chains are often found at enzyme active sites, so you might expect His 195 to be a general acid/base. Mutations drastically reduce N2 reduction. His 195 forms hydrogen bonds to sulfur S2B and bridges Fe2 and Fe3 in Figure 8, where N2 reduction is likely to occur. If His195 moves, it can form a short H bond between the imidazole N and an H bond to Fe2. If the ring rotates 180 °, no proton transfer occurs from the surface. His195 appears to be involved in the first N2 protonation event.
The Lowe and Thorneley (LT) model proposes a mechanism for dinitrogen reduction. In this model, an electron and a proton are added to the oxidized form of the enzyme (Eo) to produce E1. This is repeated three more times to form E2, E3, and E4 sequentially. Only then does N2 bind, and N2 is reduced. Two of the added electrons are accepted by H+ ions, which form H2, which is liberated on N2 binding. Hence, only 6 electrons are added to the N2 molecule, consistent with the change in oxidation numbers discussed above. The Lowe and Throneley model is shown in Figure \(\PageIndex{9}\).
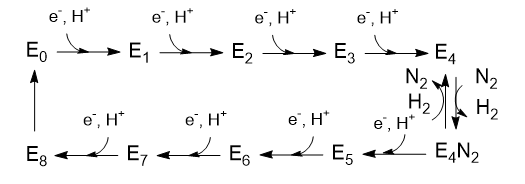
Figure \(\PageIndex{9}\): Lowe and Throneley model for electron and proton additions in nitrogenase
The crystal structure shows 2 ATP analogs bound to the reductase subunit. The reaction stoichiometry indicates that 16 ATP are used. Simple math suggests that 2 ATP are cleaved to support the entry of one electron into the complex, assuming 8 electrons are transferred (6 to N2 and 2 to protons to form H2).
Part 1 - E1-E4: A potential structure for the E4 intermediate is shown in Figure \(\PageIndex{10}\).
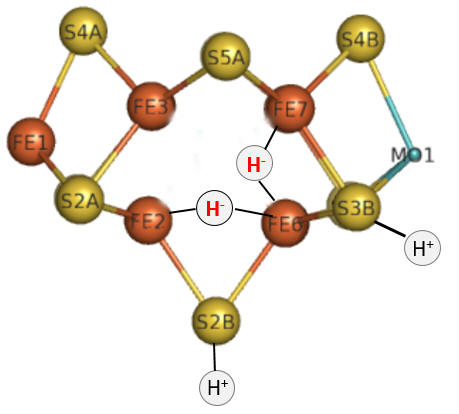
Figure \(\PageIndex{10}\): The E4 Janus intermediate in the reduction of N2.
The central carbide is not shown. This is often called the Janus intermediate because it sits halfway through the catalytic cycle. It is named after Janus, the Roman god of beginnings and transitions, and has been associated with gates, doors, doorways, and passages. Janus is typically shown with two faces, one looking to the future and one to the past (image below: DOI:10.1590/2177-6709.21.1.018-023.oin. License CC BY 4.0 Creative Commons Attribution 4.0 International)
The hydrides bridge 2 Fe ions, so these are examples of three-center, two-electron bonds. The H+ ions in Figure 10 balance the charge from the hydrides.
How does this reaction occur? We must look to organometallic chemistry to understand the mechanism and subsequent steps. Hydride equivalents have been added to the metals, associated with oxidation of the metal ions at the center. This particular reaction is called an oxidative addition. Presumably, the sulfur ions act as Lewis bases as they gain protons from a Lewis acid, probably His 195.
Oxidative addition reactions
Figure \(\PageIndex{11}\) shows oxidative additions to metal centers for three different types of reactants. L is the ligand(s), and M is the metal.

Figure \(\PageIndex{11}\): Three different types of oxidative addition reactions (after Schaller, http://employees.csbsju.edu/cschaller/ROBI1.htm)
In oxidative insertion, the metal ion's oxidation state increases, hence the name oxidative. The hydrogens are now hydrides. Note the H2 insertion example, which is similar to the proposed hydride additions to the M cluster. Oxidative reduction occurs most readily when the metal ion's two oxidation states are stable. It is likewise favored for metal centers that are not sterically hindered (makes sense if A-B is to be added) and if A-B has a low bond dissociation energy.
One way to study reaction intermediates is to trap them in place. If N2 can't access the binding site and the temperature is reduced, the accumulated hydrides (and for charge balance, the H+s) in E4 might leave in the opposite reaction, reductive elimination. We will discuss this below, as the reaction goes back to E1. In the elimination, they can form H2 as the metal gains back electrons in a reduction. The Val70Ile discussed above would allow trapping of an intermediate.
Nitrogenase Reaction: Part 2 - Reduction of N2
Steps E4 to E5 seem a bit bizarre as H2 gas is released. This would seem to waste ATP, but we should trust that evolution has led to this mechanism for a reason. This mechanism, the reverse of oxidative addition, is another classic organometallic reaction: reductive elimination.
Reductive elimination reactions
In this reaction, a molecule is eliminated or expelled from the complex as the metal ion is reduced, gaining two electrons. Figure \(\PageIndex{12}\) shows reductive elimination. Reductive elimination occurs most readily at higher-oxidation-state metal centers, which are stabilized upon reduction. It occurs most readily with electron-rich ligands and when the other surrounding ligands are bulky. The dissociating species must also be cis to each other in the transition-metal complex so they can bond to each other when they leave.
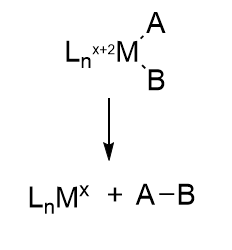
Figure \(\PageIndex{12}\): Reductive Elimination reaction (after Schaller, ibid)
Oxidative addition and reductive elimination (OA/RE) reactions at metal centers are often coupled together in organometallic catalytic cycles, much like a histidine can act as a general acid and then accept a proton back as a general base to complete the catalytic cycle. In the OA/RE reactions at metal centers, some rearrangements or other modifications can also occur. Think about it. The FeS clusters must return to their original oxidation state after the complete LT cycle. We will encounter another organometallic reaction after the addition of N2, migratory insertion, in the second half of the reaction. Another advantage of coupling OA/RE is that the positive charge, or oxidation state, on the transition-metal complex does not become too high, which would be unstable. Making a cation more positive becomes more difficult, much as removing a second proton from a polyprotic acid is more difficult than removing the first (as reflected in the higher pKa for removing the second proton).
Is H2(g) released? To study this, investigators have used alternative substrates such as acetylene, HC≡CH (similar to N≡N), in aqueous systems with D2 and N2. It is helpful to review Figure \(\PageIndex{9}\) again.
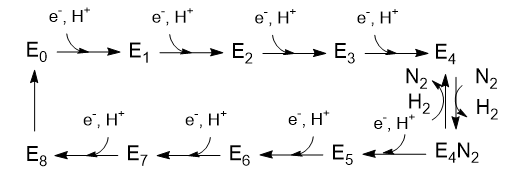
Figure \(\PageIndex{9}\): Lowe and Throneley model for electron and proton additions in nitrogenase
Acetylene was reduced to form C2H2D2 and C2H3D. Therefore, E4 must have had two Ds, and E2 probably had one. These results support the reversible reductive elimination mechanism for the E4 to E4:N2 reaction above. Previously, it had been shown that H+ is reduced by D2 in the presence of D2 and N2 in an aqueous system, so these results are consistent. In addition, deuterium from D2 is not incorporated into products (C2H2D2, C2H3D, or HD) in the absence of N2.
Let's return for a moment to the bridging hydrides as shown in Figure \(\PageIndex{10}\) again.
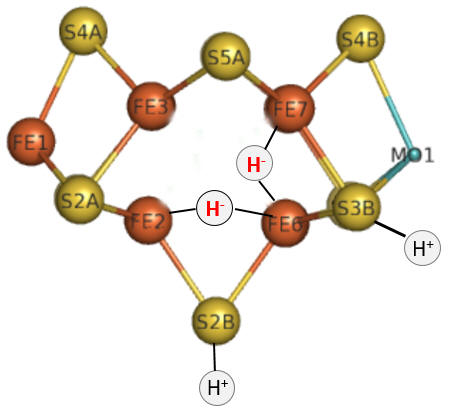
Figure \(\PageIndex{10}\): The E4 Janus intermediate in the reduction of N2
The reductive elimination of the two proximal bridging hydrides likely forms H2. The bridging hydrides, which are strong bases, are much less likely to be protonated than terminal hydrides. A simple competing protonation reaction could also form H2; if it did, the bridging hydrides would lose their reducing equivalents. Hence, the bridging hydrides (shared by two metal centers) are more stable and can "wait" for the incoming N2 reactants before losing their reducing equivalents. They may convert to terminal hydrides eventually to facilitate substrate reduction (hydrogenation).
The Janus intermediate E4 is now positioned to bind N2 and release H2. For each H- that binds to the M cluster, two H+s bind to the M cluster sulfides for electrostatic stabilization.
Migratory Insertions
We need to consider one last common reaction at a metal center: migratory insertions (MI). In an MI reaction, a group attached to a metal center transfers to another group attached to the same metal. Figure \(\PageIndex{13}\) shows four examples of MI reactions.
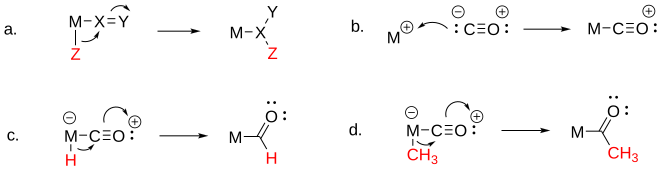
Figure \(\PageIndex{13}\): Migratory insertion reactions at metal centers
Panel (a) shows a generic MI reaction. Panel (b) shows the interaction of carbon monoxide, :C≡O: (isoelectronic and analogous to :N≡N:), with a metal center. Panel (c) shows how a hydride could engage in 1:1 insertion as it shifts and covalently bonds to the first atom of another ligand bound to the metal center. Note that the metal does not have to have a negative charge. This could theoretically be important for reducing N2 bound as a ligand through a coordinate covalent (dative) bond to the metal. Panel (d) shows alkyl group migration.
We will see that the MI reaction involves adding Hs to N2, starting not with N2 but with the N2H2 stage, as the intermediates insert into an Fe-H bond. N2 is not reactive to hydride insertion, as carbon monoxide (CO) is, which provides a positive oxygen to facilitate electron flow during insertion. In addition, the oxygen becomes neutral after the reaction.
This provides a compelling explanation for how natural selection favored the FeMo cluster for nitrogen fixation. The interaction of two hydrides requires a 4 Fe face (coordinated with the carbon) that allows for the storage of reducing equivalents for the initial reduction of N2. The large M cluster is more stable and held together more effectively by a central carbide anion. This also keeps the metal centers from changing their oxidation state by more than one charge unit.
The protons that form N2H2 come directly from the two H2 attached to the two sulfurs. They can't come from the hydrides, which are released as H2. The seemingly wasteful reductive elimination of H2 and energy is required to allow the kinetically unreactive N2 molecule to bind to the reduced and activated 4Fe face, which is also electrostatically facilitated by the two bound protons in what has been called a push (reductive)-pull (protonation) reaction. This first step in N2 reduction to N2H2 is the hardest.
Oxidation States of Nitrogenase Fe Centers
First Half: Assigning specific oxidation states to each Fe ion in the M complex is difficult. Instead, we can assign relative changes in the oxidation states as the reaction proceeds from E0 to E4. In each step of the LT model, 1 electron is added. We will first assign this to an average Fe ion, M0, with an arbitrarily assigned oxidation state of 0. After adding 1 electron, the oxidation state would change from M0 to M-1 as the metal is reduced. The M-1 state is then oxidized as an electron is transferred to H+, and when 2 electrons are transferred, a single H- is formed. The diagram in Figure \(\PageIndex{14}\) illustrates the change in oxidation state as the system transitions from E0 to E4.

Figure \(\PageIndex{14}\): Change in oxidation state in going from E0 to E4
The red boxes highlight thermodynamic-cycle-like steps that show how changes in the redox state of the Fe ions (M) can be visualized. Note that, going from E0 to E4, the actual oxidation state of M alternates between 0 and +1, then back to 0. That is quite amazing given that 4 electrons have been added. Note also that in the red box from step E0 to E1, M changes from -1 to +1, which corresponds to our description of an oxidative addition when the metal center loses two electrons. This mechanism suggests that nitrogenase can be considered a "hydride storage device."
Second Half (facing forward to production NH3):
How does N2 initially interact with E4? It depends on how the hydrides are released as H2, as evidence shows it occurs via reductive elimination (re), not hydride protonation (hp). On addition, the N2 is very quickly converted to diazene, HN=NH, with the departing H2 taking with it 2 H+s and 2 electrons (or reducing equivalents). These events could occur as shown in Figure \(\PageIndex{15}\).

Figure \(\PageIndex{15}\): Reaction mechanism for the formation of N2H2
Now, with N2 bound as diazene (N2H2) and H2 released, the remaining reaction could occur as shown below. One new step, a migratory insertion, is shown in Figure \(\PageIndex{16}\).

Figure \(\PageIndex{16}\): Reaction mechanism for the conversion N2H2 to NH3.
The two halves of the reaction are similar, using bridging hydrides; the E4 Janus intermediate links the two halves.
Summary
(Summary written by Claude, Sonnet 4.6, Anthropic)
This chapter describes the biological nitrogen cycle and the enzyme nitrogenase, which catalyzes the most chemically demanding reaction in microbial biochemistry — the reduction of atmospheric N₂ to NH₄⁺ — examining the thermodynamic and kinetic basis for N₂ inertness, the remarkable metal cluster architecture of nitrogenase, and the organometallic reaction mechanisms that accomplish this transformation under ambient biological conditions.
Nitrogen occupies a central position in biochemistry: all four major biomolecule classes (lipids, carbohydrates, nucleic acids, and proteins) contain nitrogen, and all depend ultimately on its bioavailability. Atmospheric N₂ (80% of air) is chemically inert under ambient conditions because of its N≡N triple bond (bond dissociation energy ~945 kJ/mol) and its lack of a dipole moment, creating a prohibitively high activation energy barrier. The biological nitrogen cycle encompasses five transformations that collectively maintain bioavailable nitrogen: fixation (N₂ → NH₄⁺ by nitrogenase in prokaryotes); nitrification (NH₄⁺ → NO₂⁻ → NO₃⁻ by aerobic ammonia-oxidizing and nitrite-oxidizing bacteria via hydroxylamine intermediates); denitrification (NO₃⁻ → N₂ by anaerobic bacteria); anammox (NO₂⁻ + NH₄⁺ → N₂ by specialized anaerobic bacteria); and ammonification (organic nitrogen → NH₄⁺ by decomposition). Human industrial nitrogen fixation via the Haber-Bosch process now contributes more fixed nitrogen than the entire biosphere, with excess NH₄⁺ from agricultural runoff overwhelming natural nitrification/denitrification balancing, acidifying waterways, causing eutrophication, and generating N₂O — a greenhouse gas approximately 300 times more potent than CO₂.
The Haber-Bosch reaction (N₂ + 3H₂ → 2NH₃) is thermodynamically favorable at room temperature (ΔG° = −16.4 kJ/mol; ΔH° = −46.2 kJ/mol) but kinetically blocked. Because the reaction is exothermic, raising the temperature to overcome the activation energy paradoxically disfavors product formation by shifting Keq. The industrial solution — high pressure (200–400 atm) to shift equilibrium toward two product molecules from four reactants, moderate-high temperature (400–500°C) for kinetic feasibility, and a magnetite/metal oxide catalyst — exemplifies thermodynamic-kinetic compromise but requires ~1–2% of global energy production. Biological nitrogen fixation by nitrogenase solves the same problem under ambient pressure and temperature at extraordinary energetic cost — 16 ATP per N₂ reduced — using a sophisticated metalloprotein architecture.
Nitrogenase is a two-component holoenzyme. The Fe-protein (reductase, Av2 in Azotobacter vinelandii) is a homodimer (E/F subunits) containing a single 4Fe-4S F cluster, binding sites for ferredoxin or flavodoxin (the electron donors), and two ATP-binding sites. Electron entry from the reduced carrier protein triggers ATP hydrolysis — two ATPs per electron transferred — which drives conformational changes facilitating dissociation and reassociation of the Fe-protein from the MoFe-protein, a cycle essential for each electron delivery. The MoFe-protein (diazotrophase, Av1) is an α₂β₂ heterotetramer. The 8Fe-7S P cluster, located at the α/β interface, mediates electron transfer from the F cluster to the M cluster. The FeMo-cofactor M cluster (Fe₇MoS₉C) resides in the α subunit and is the site of N₂ binding and reduction. Mo is coordinated to three sulfides and the bidentate oxygen ligands of 3-hydroxy-3-carboxy-adipic acid. Critically, an interstitial carbide ion (derived from the methyl group of S-adenosylmethionine during cofactor assembly) occupies the center of the M cluster, where it stabilizes the Fe₇ cage without being incorporated into products during turnover. Val70 acts as a gatekeeper controlling substrate access to the M cluster; His195, hydrogen-bonded to sulfur S2B and bridging Fe2/Fe3, likely acts as a general acid/base facilitating the first N₂ protonation event. Nitrogenase is irreversibly destroyed by O₂, requiring nitrogen-fixing organisms to employ anaerobic niches, heterocysts with thickened cell walls, or leghemoglobin buffering (in root nodule symbioses with legumes) to maintain local anaerobiosis.
The catalytic mechanism follows the Lowe-Thorneley (LT) kinetic model, in which the enzyme cycles through states E₀ to E₄ before N₂ binds, and then E₄ to E₈ to complete reduction. In the first half (E₀→E₄), each step delivers one electron and one proton to the M cluster. However, rather than directly reducing N₂, the first four electrons are used to produce two bridging hydrides (Fe-H-Fe) on the iron face of the M cluster through oxidative addition — analogous to H₂ oxidative insertion into transition metal complexes, where the Fe oxidation state increases as H₂ is split into two hydride ligands. Bridging hydrides (three-center, two-electron bonds shared between two iron centers) are much more stable than terminal hydrides toward protonation and can serve as a thermodynamic reservoir of reducing equivalents, waiting for the arrival of N₂ without premature loss as H₂. Two H⁺ ions are simultaneously bound to the M cluster sulfides for electrostatic charge stabilization. This accumulation defines the E₄ "Janus intermediate" — so named because it stands at the mechanistic midpoint, looking backward toward hydride accumulation and forward toward N₂ reduction.
The central, seemingly wasteful step — reductive elimination of H₂ from E₄ upon N₂ binding — is actually thermodynamically essential. Reductive elimination (the microscopic reverse of oxidative addition) releases H₂ as the Fe center gains two electrons, creating a highly reduced, low-valent 4Fe face uniquely capable of binding and activating the kinetically inert N₂ molecule. This reductive-pull is coupled with a proton-push from the two sulfide-bound H⁺ ions, generating diazene (HN=NH) from the bound N₂ in the first bond-breaking step. Experimental evidence supports this mechanism: when D₂ is used in place of H₂ in the presence of the alternative substrate acetylene (HC≡CH, isoelectronic with N₂), deuterium is incorporated into reduced acetylene products (C₂H₂D₂, C₂H₃D), confirming that the hydrides in E₄ are exchangeable and that reductive elimination is reversible.
The second half of the reaction (E₄:N₂→2NH₃) proceeds through sequential migratory insertion reactions — a third class of organometallic mechanism in which a ligand bound to the metal migrates to insert into a neighboring M-H bond. Diazene (HN=NH) inserts into Fe-H bonds, with the metal center transferring hydrides to successive nitrogen intermediates — diazenido (HN=N), hydrazido (H₂N-N), hydrazine (H₂N-NH₂), and ultimately two NH₃ molecules. Throughout this sequence, the oxidation state of individual Fe centers changes by no more than ±1 unit per step, an elegant constraint that maintains cluster stability and prevents irreversible over-oxidation of the reactive metal centers. The complete catalytic cycle requires 8 electrons, 8 H⁺, and 16 ATP per N₂: 6 electrons reduce N₂ to 2NH₃, and 2 electrons plus 2 H⁺ produce the obligate H₂ byproduct. This remarkable reaction — accomplishing under ambient conditions what industrial chemistry requires extreme pressure, temperature, and metal catalysis to achieve — stands alongside the oxygen-evolving complex of photosystem II as one of the most chemically sophisticated and biologically consequential enzymatic achievements in nature.